End-to-end GROMACS 2024.4 GPU-accelerated MD pipeline — from complex preparation and equilibration to production runs, binding energy calculations, and publication-quality analysis.
Our MD simulation platform provides a complete workflow built on GROMACS 2024.4 with GPU acceleration. The pipeline covers every step from receptor-ligand complex preparation with the CHARMM36 force field, through energy minimization and NVT/NPT equilibration, to production MD simulations.
Advanced post-simulation analysis includes MM-PBSA/GBSA binding free energy calculations, PCA analysis for conformational dynamics, and a comprehensive suite of publication-ready visualization tools for RMSD, RMSF, SASA, Rg, hydrogen bonds, FEL, DCCM, and more.
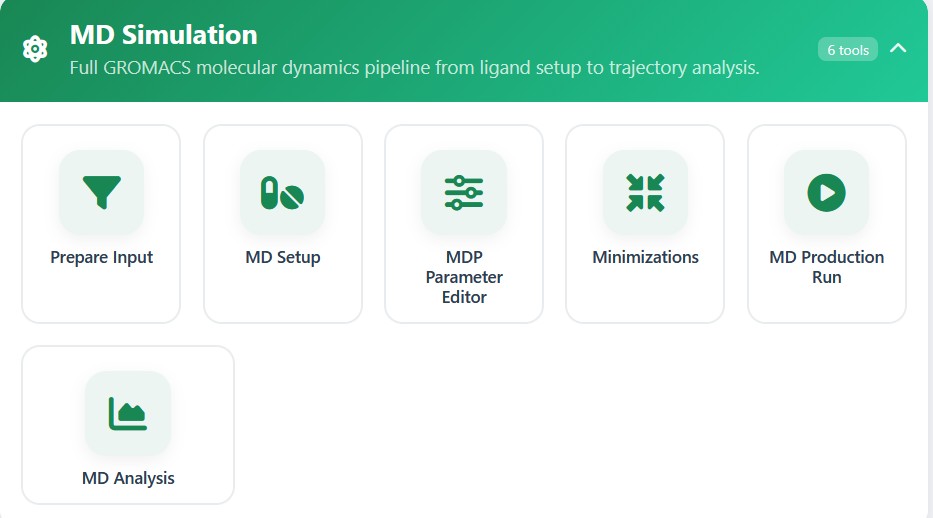
A complete 6-step pipeline from preparation to analysis
Ligand topology with SwissParam / CGenFF
REC-LIG complex with CHARMM36 force field
EM → NVT → NPT with MDP editor
GPU-accelerated simulation runs
Binding free energy calculations
Plots, PCA, FEL, DCCM
Six integrated steps in the web-based GROMACS 2024.4 MD platform
Real outputs from the MD simulation platform
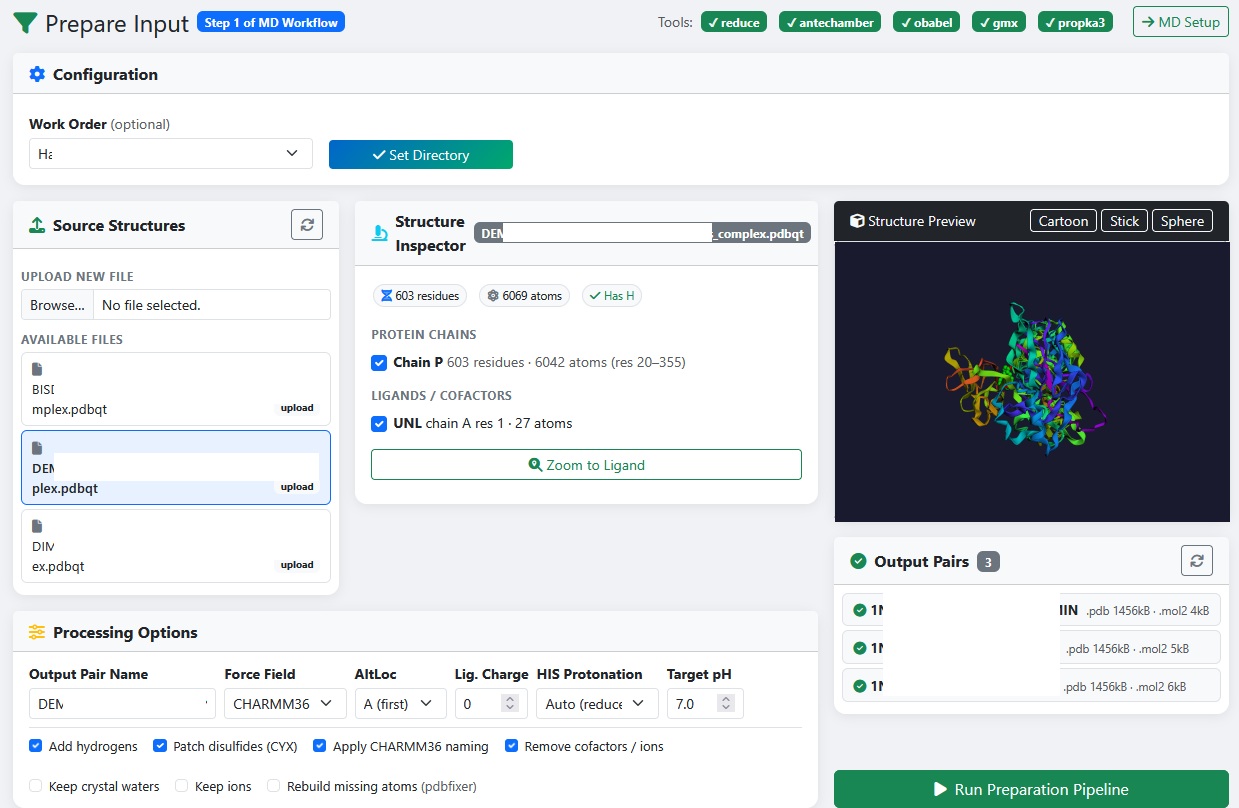
Upload receptor-ligand complexes, inspect chains/atoms, configure CHARMM36 force field, protonation (PROPKA3), and run preparation pipeline
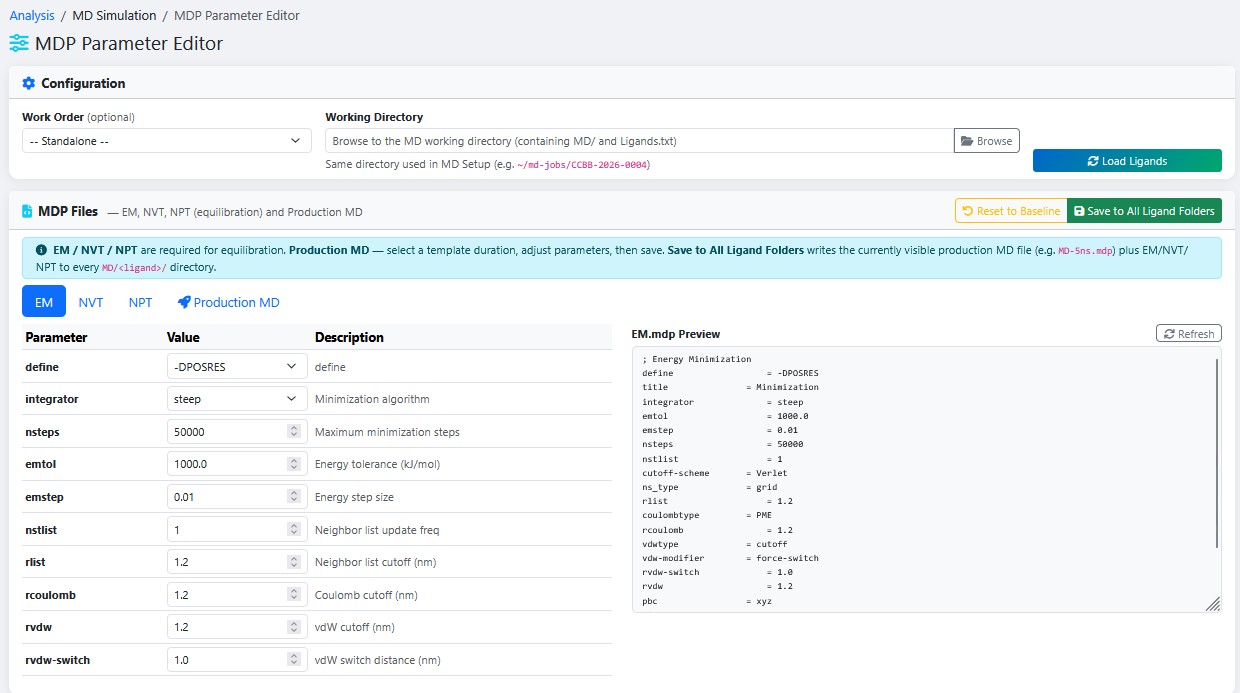
Web-based editor for EM, NVT, NPT, and Production MD parameters with live preview and batch save to ligand folders
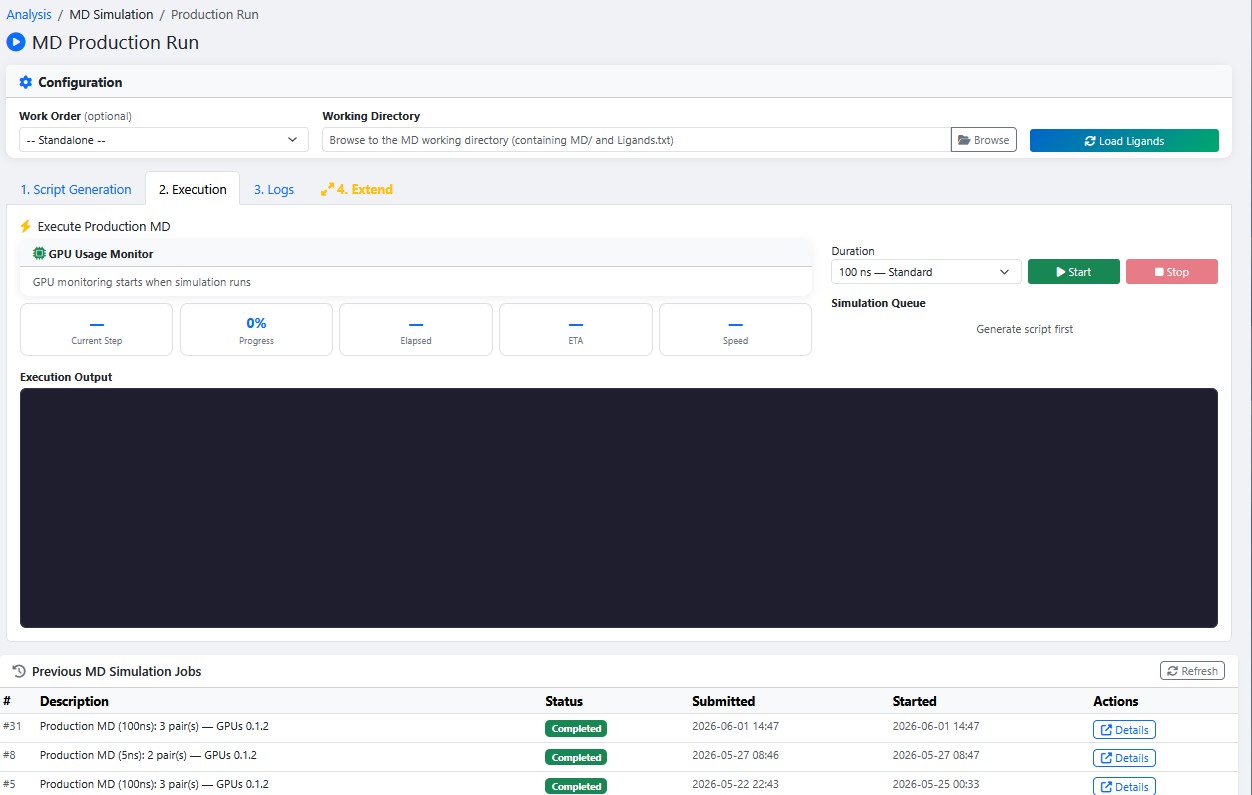
GPU usage monitoring, real-time simulation progress, duration selection (5 ns – 200 ns), and completed job history
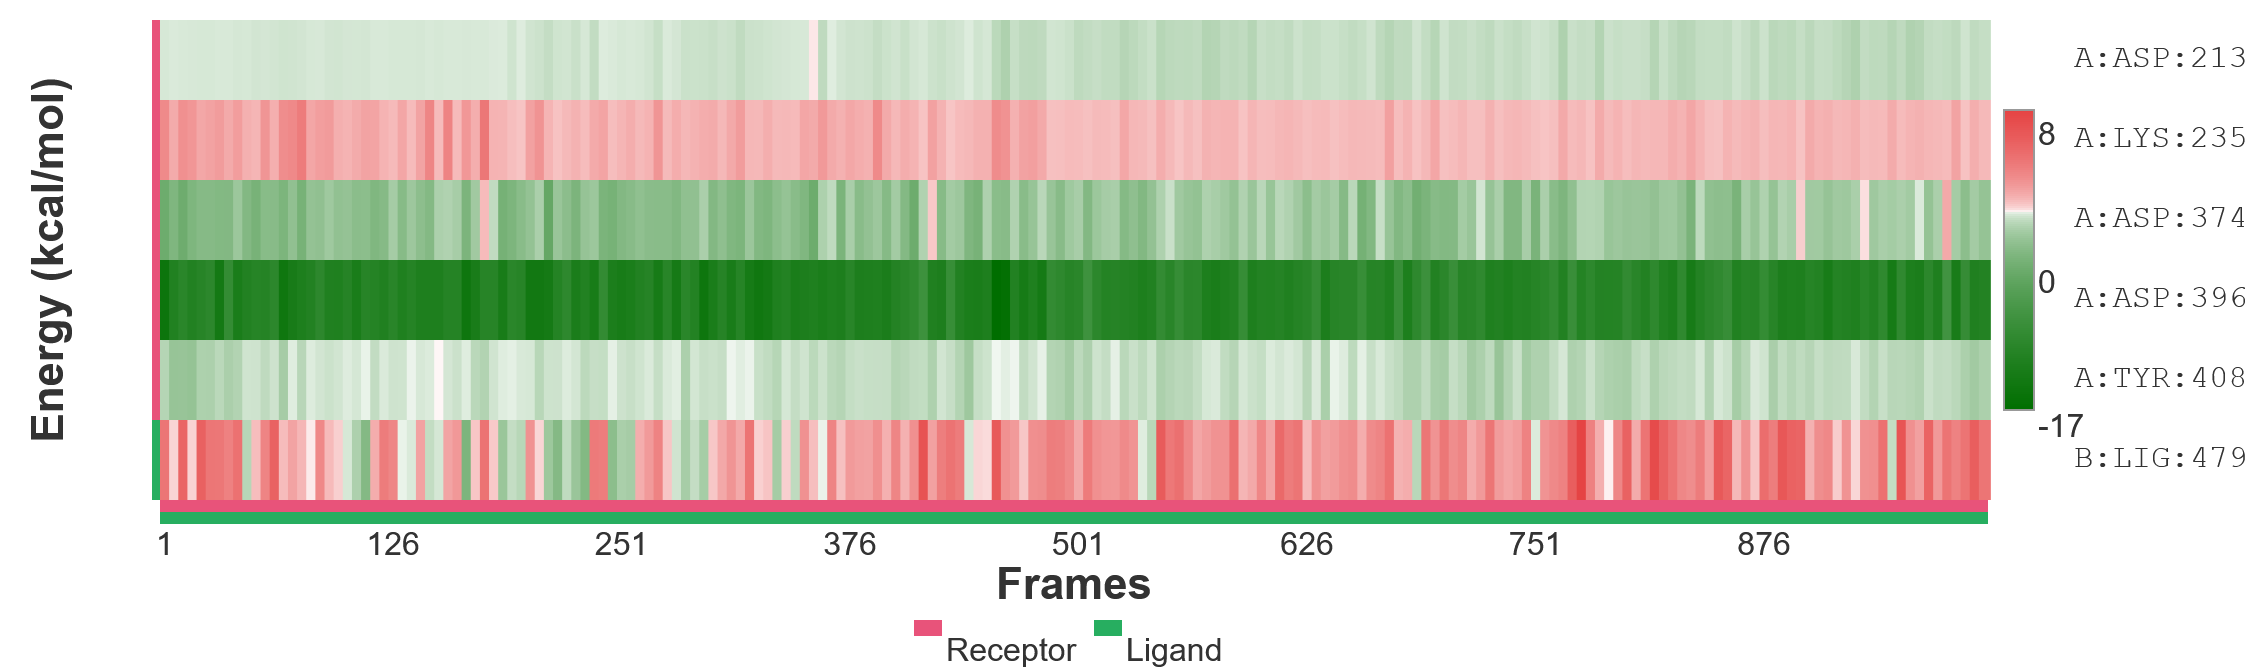
Per-residue energy contribution heatmap showing electrostatic, van der Waals, and solvation terms
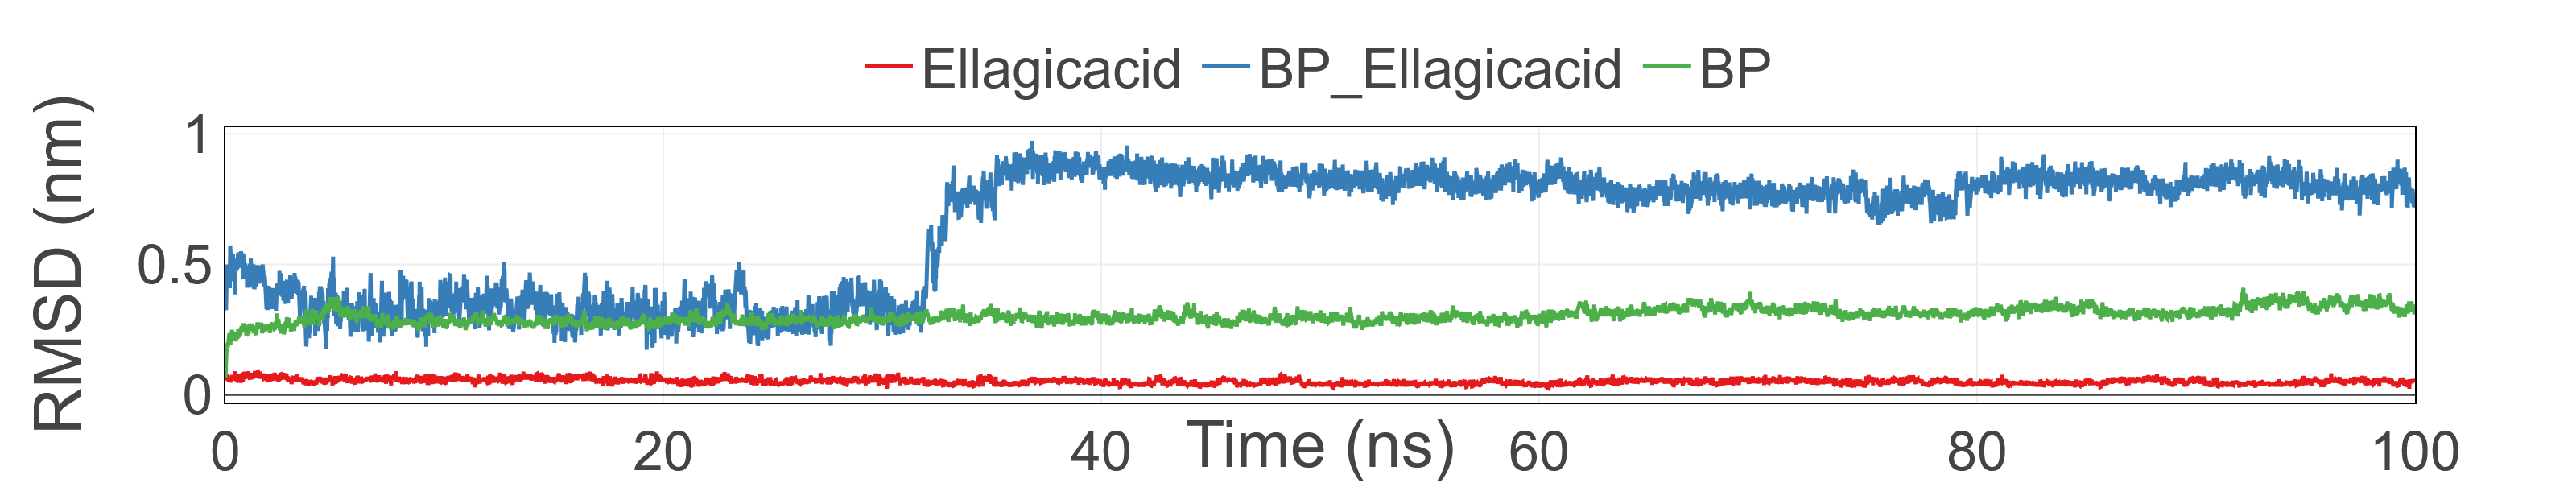
Root Mean Square Deviation over simulation time for structural stability assessment
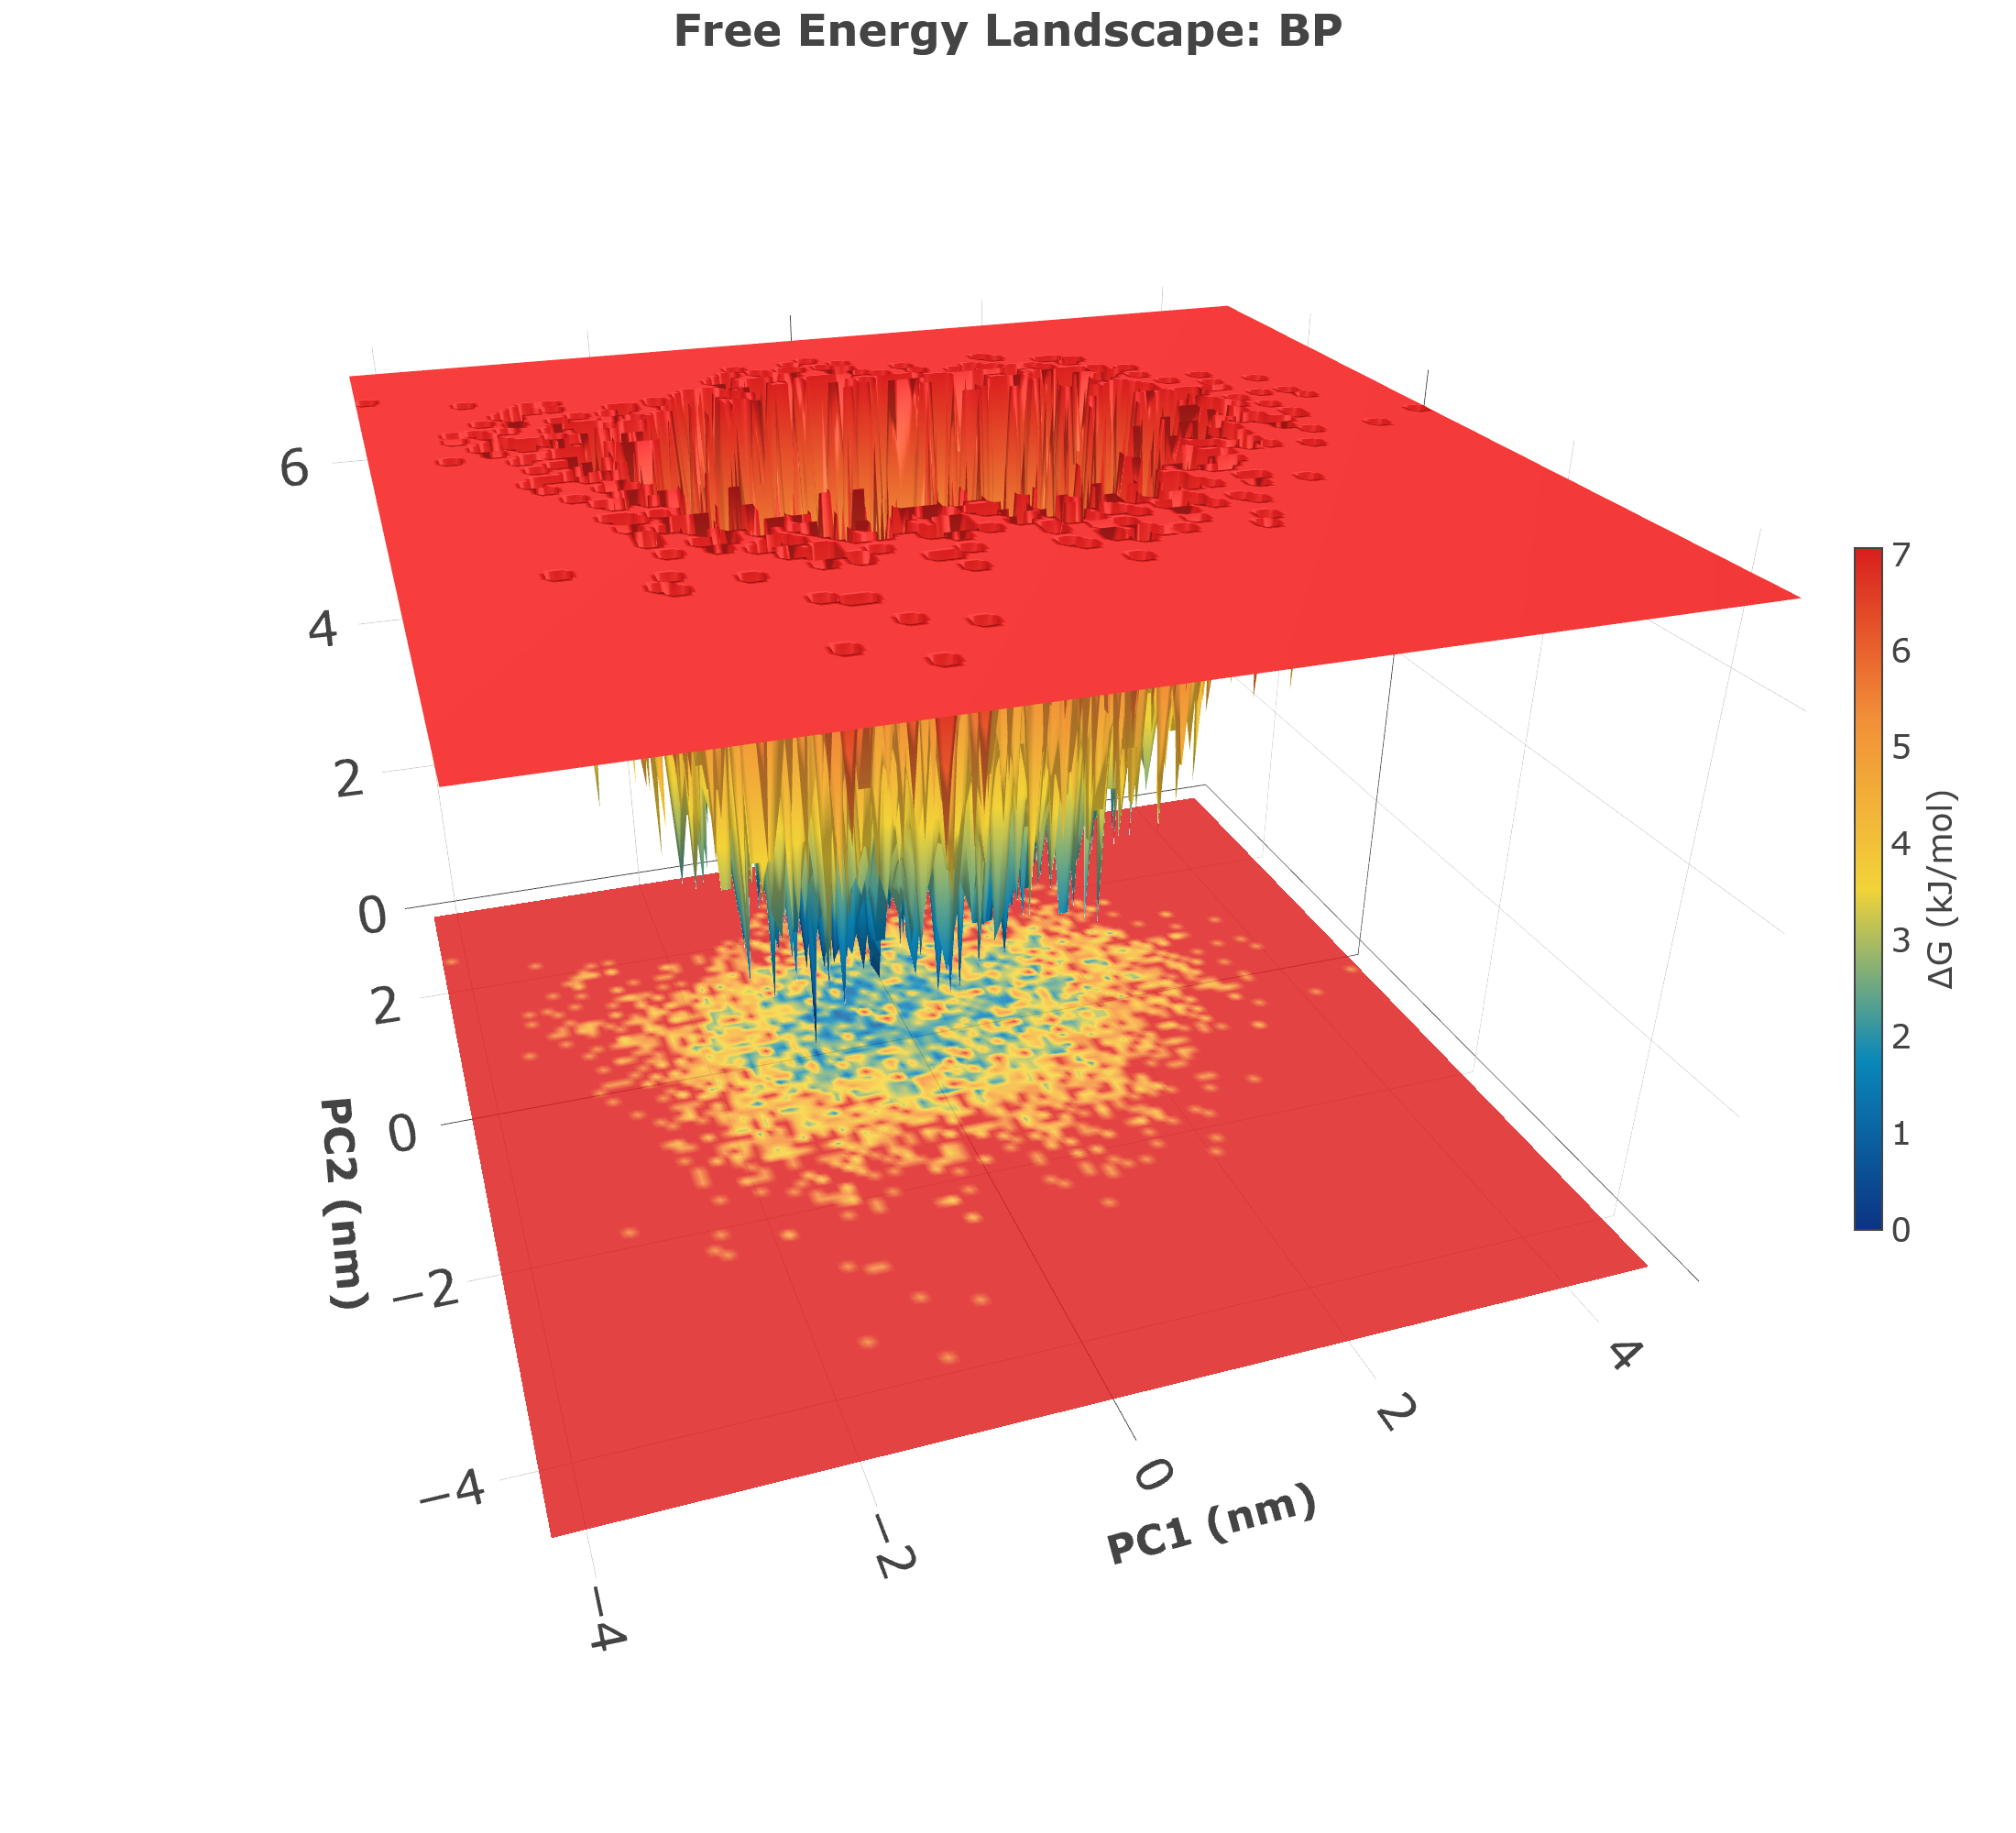
FEL surface with contour projection and conformational energy minima identification
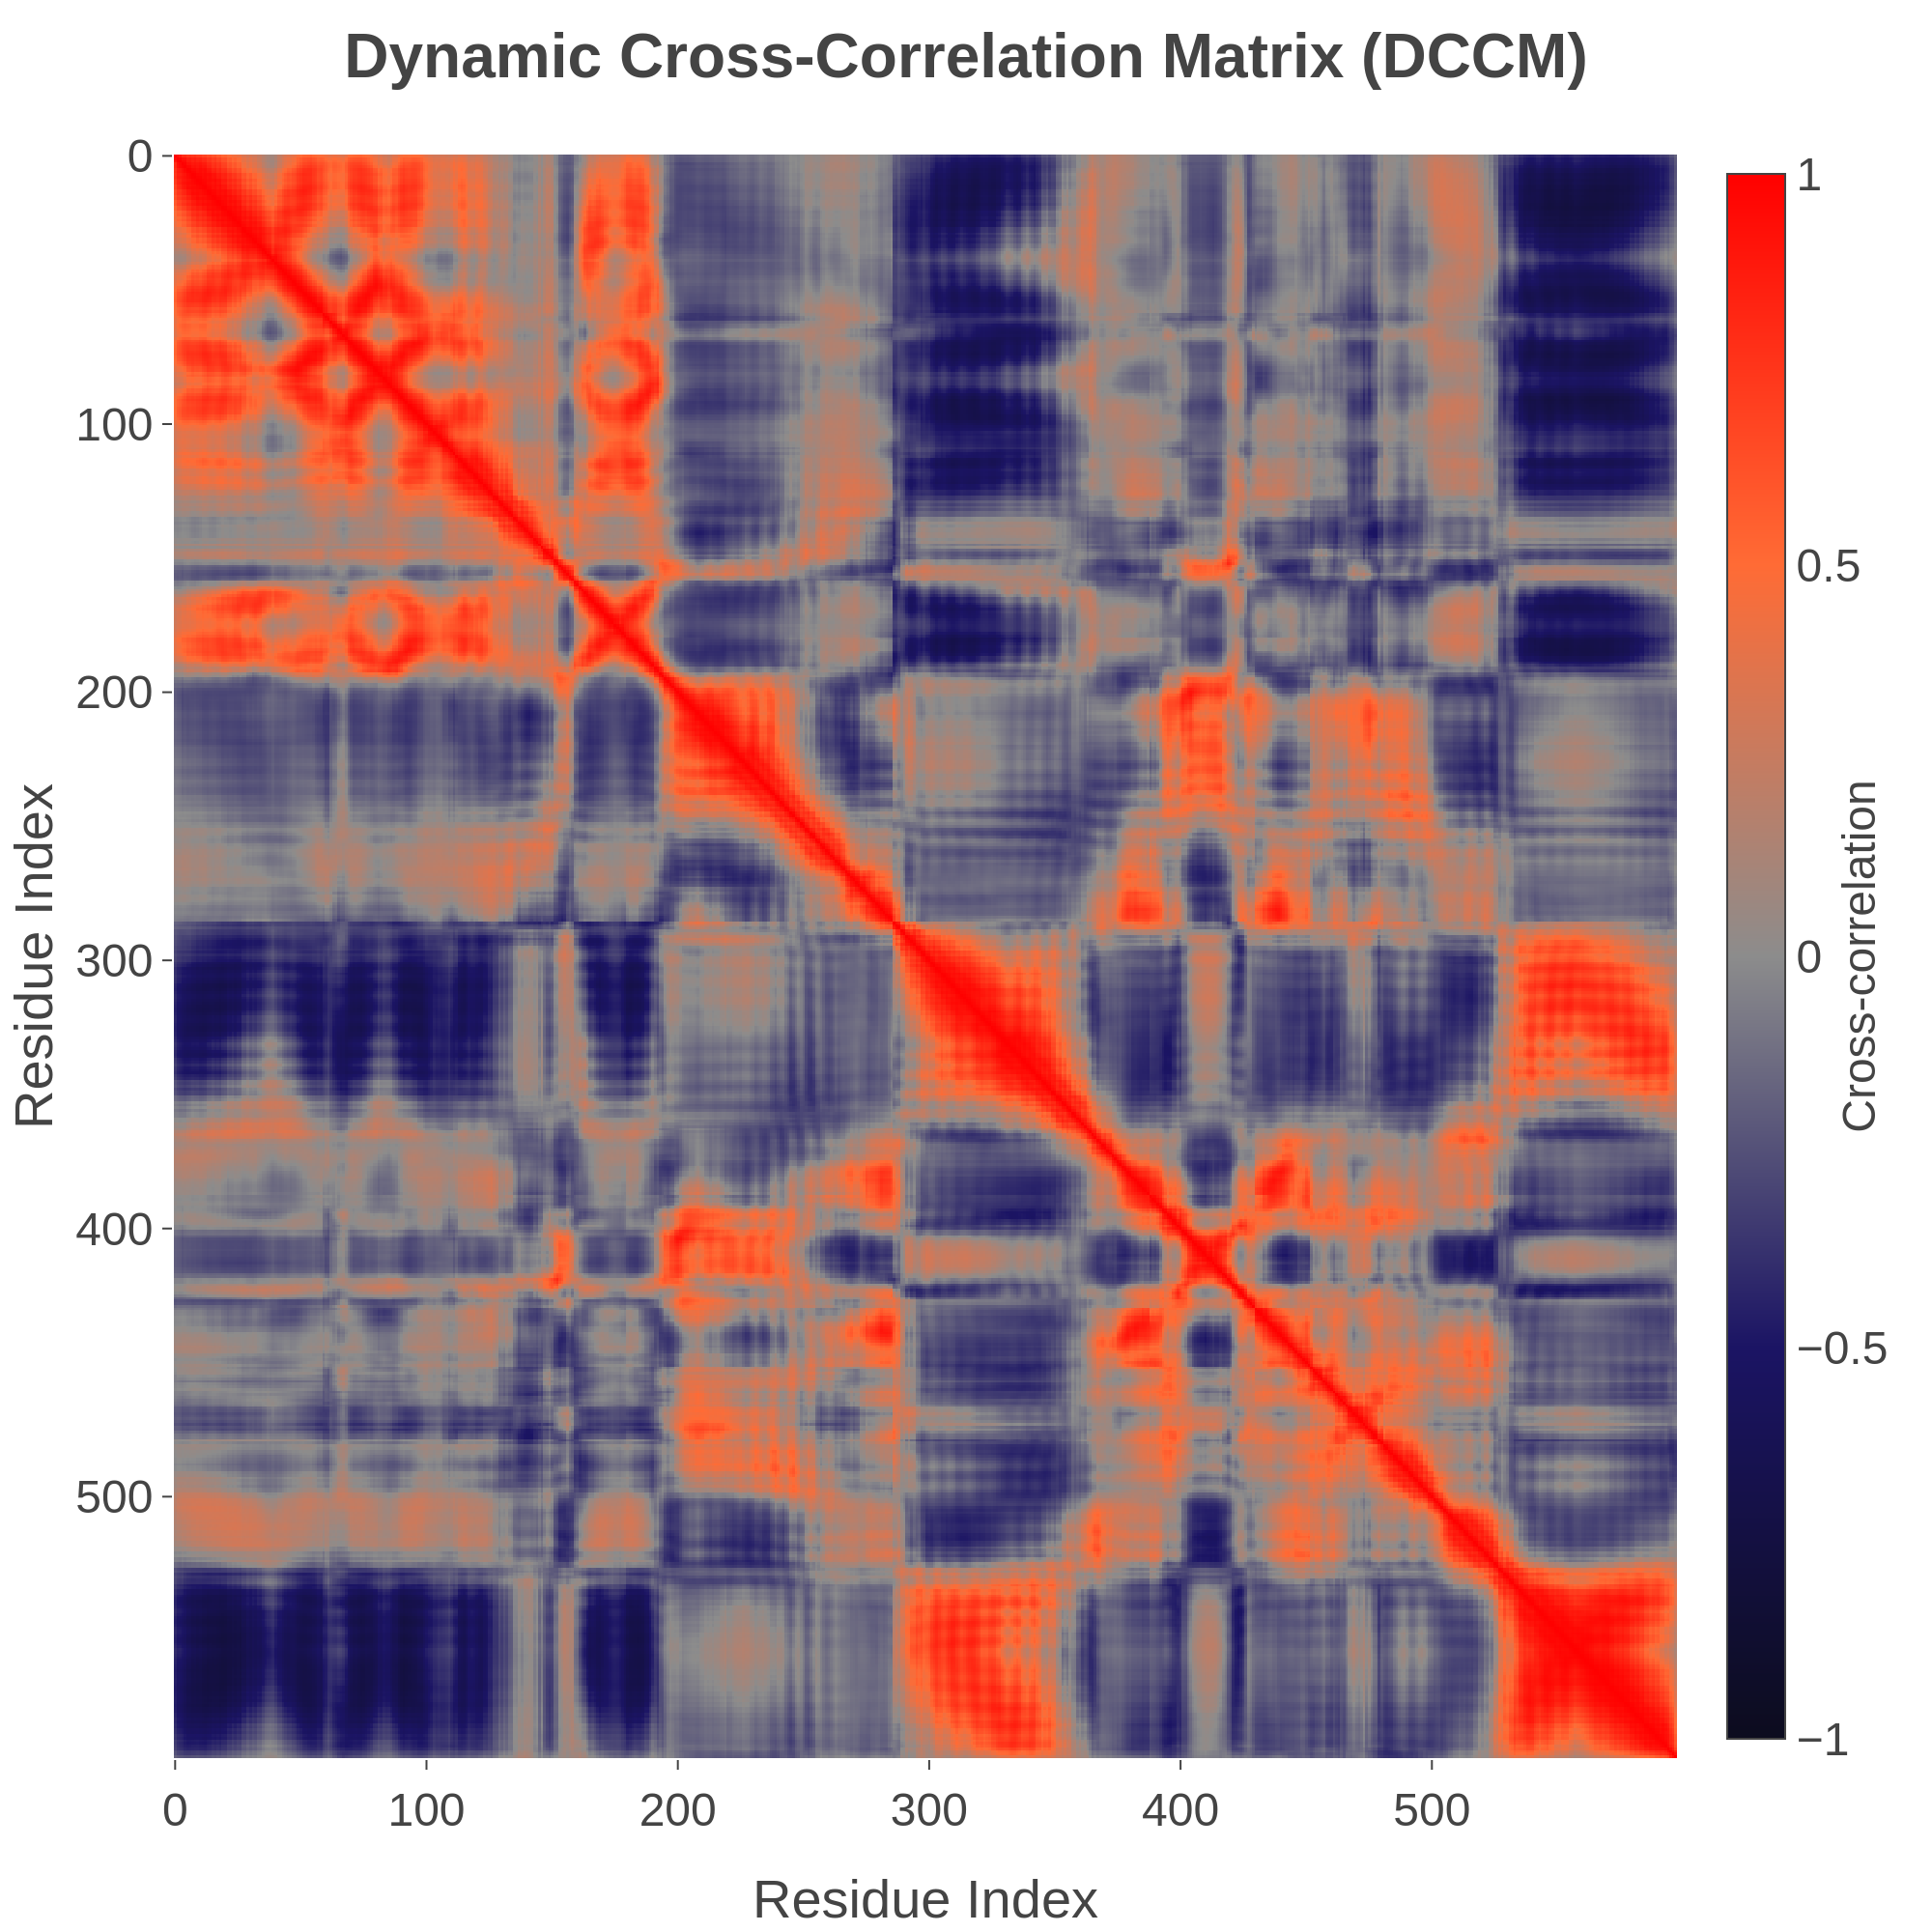
Residue-residue dynamic cross-correlation analysis with customizable color scale
Watch a complete MD simulation workflow
From complex preparation to publication-ready analysis — Coming soon
Sign up to access the full GPU-accelerated MD simulation platform
Get Started Contact Us