What we do
Preparation
Receptor & Ligand Preparation
- Structures from the PDB by ID, or your own models (homology, AlphaFold)
- Choice of chains and co-crystallised ligands; waters removed; hydrogens and charges added
- Docking box around a bound ligand, chosen residues or the whole protein; AutoGrid4 / AGFR grid maps
- Ligands from SDF / MOL2 / PDB, PubChem (names, SMILES, CIDs) or virtual-screening hits; 3D coordinates and MMFF94 minimisation
- Peptide ligands built from their sequence
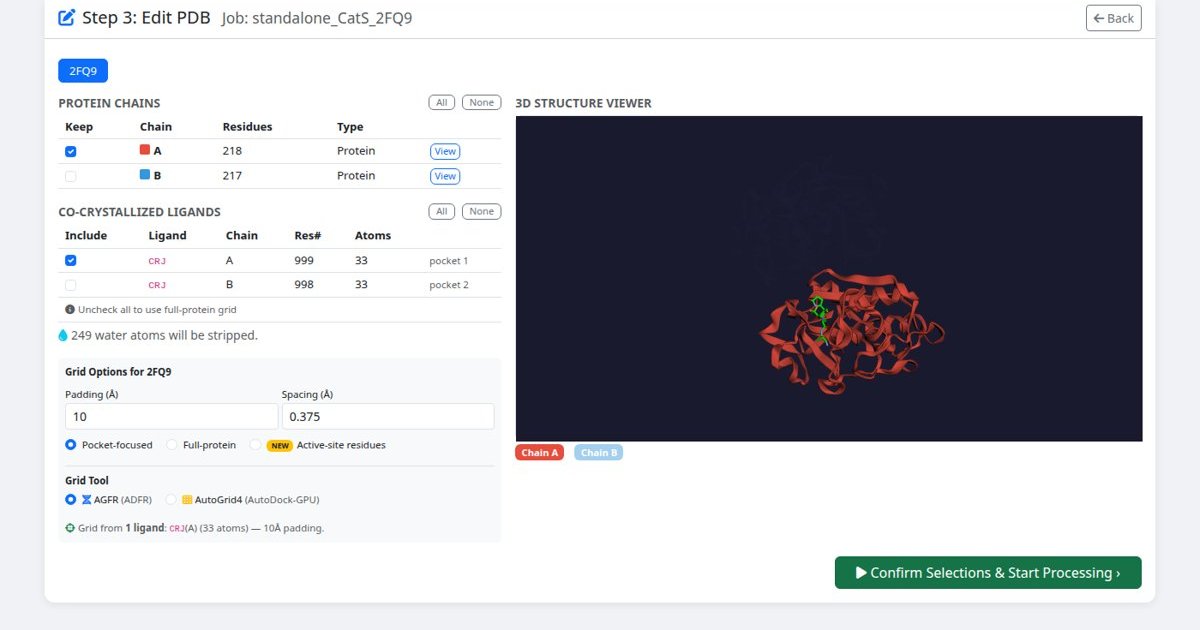
Receptor editor for cathepsin S (PDB 2FQ9): chains, the bound inhibitor and the grid options.
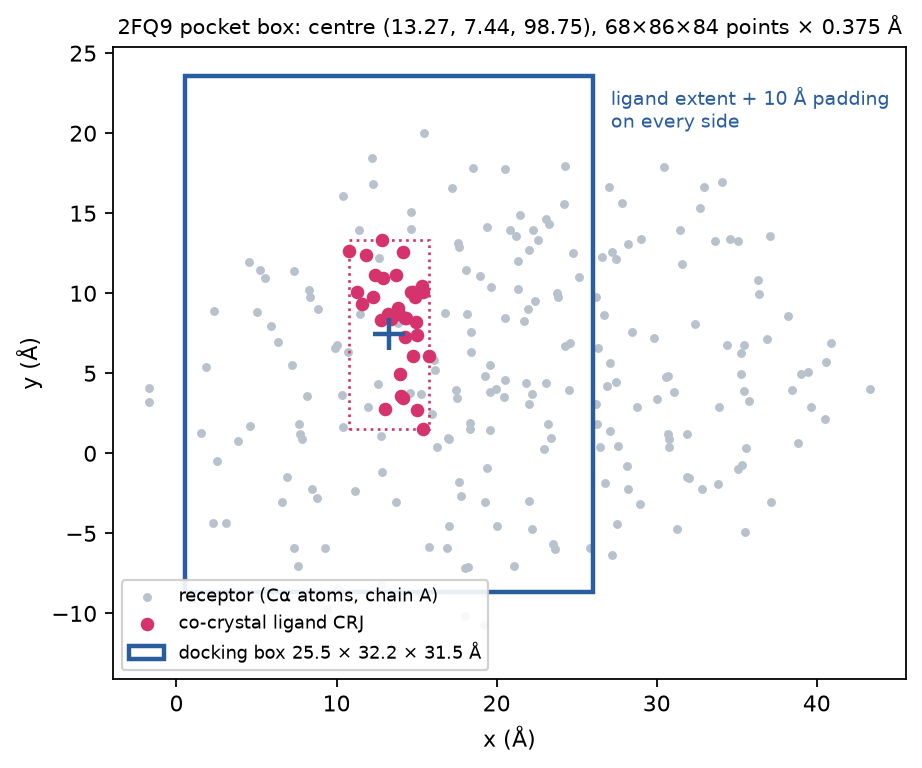
The docking box: ligand extent plus 10 Å.
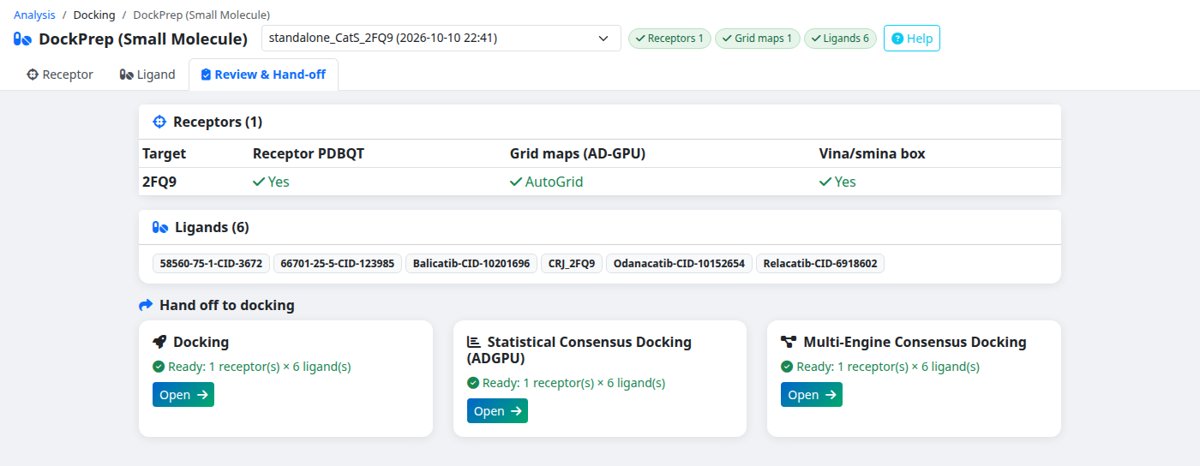
Readiness check before docking, for each engine.
Docking
Three Docking Engines
- AutoDock-GPU — AutoDock4 scoring, many independent runs per ligand clustered into binding modes, on three NVIDIA A30 GPUs
- AutoDock Vina and smina — empirical scoring on the CPUs, reproducible with a recorded seed
- All engines search the same box, so results can be compared and combined
- From a handful of ligands to large libraries
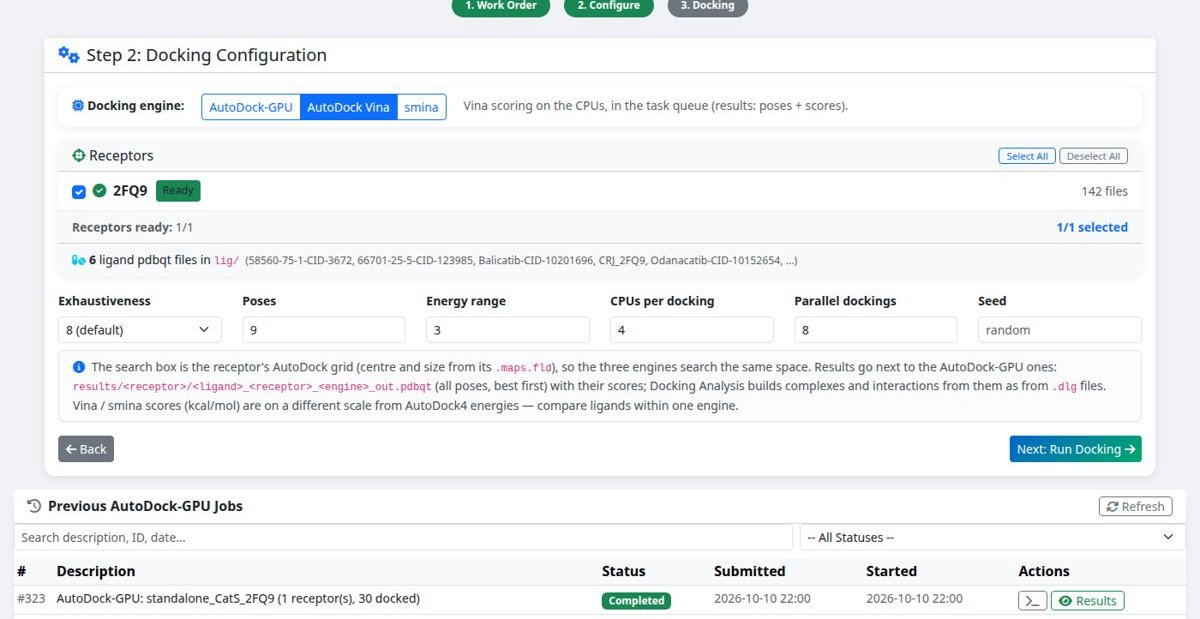
Choosing the engine and its settings for a docking job.
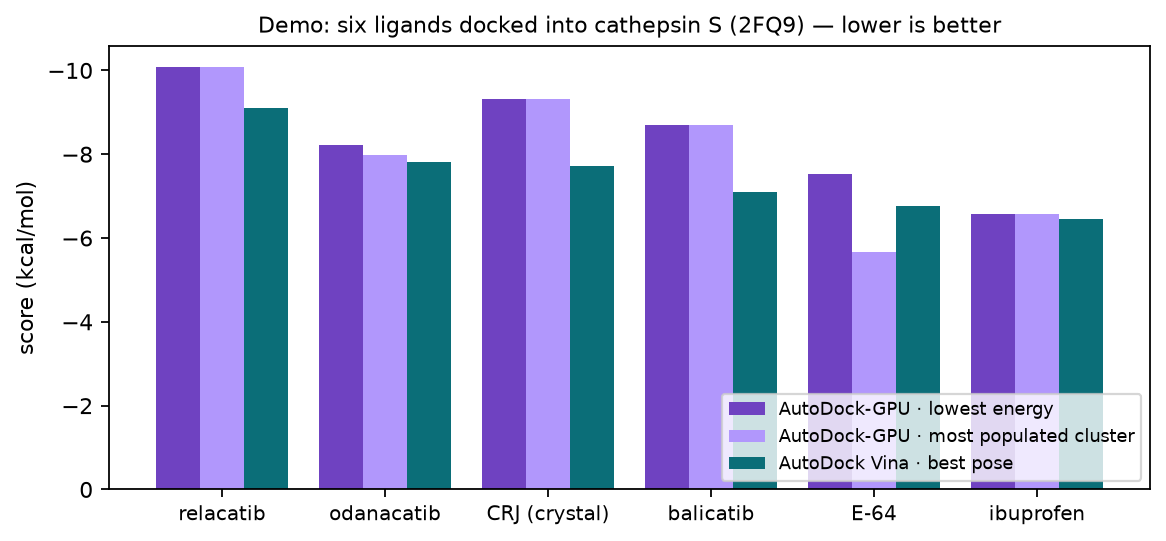
Six ligands docked into cathepsin S with AutoDock-GPU and AutoDock Vina.
Analysis
Poses, Interactions & Reports
- Pose selection by cluster population or lowest energy
- Protein–ligand complexes for every ligand and engine
- Interactions with PLIP: hydrogen bonds, hydrophobic contacts, π-stacking, salt bridges, halogen bonds, water bridges
- 3D images of the complex and the pocket, 2D interaction diagrams, interaction tables
- Excel summaries and a publication-style PDF report

Relacatib in the cathepsin S pocket: 3D view and 2D interaction diagram.
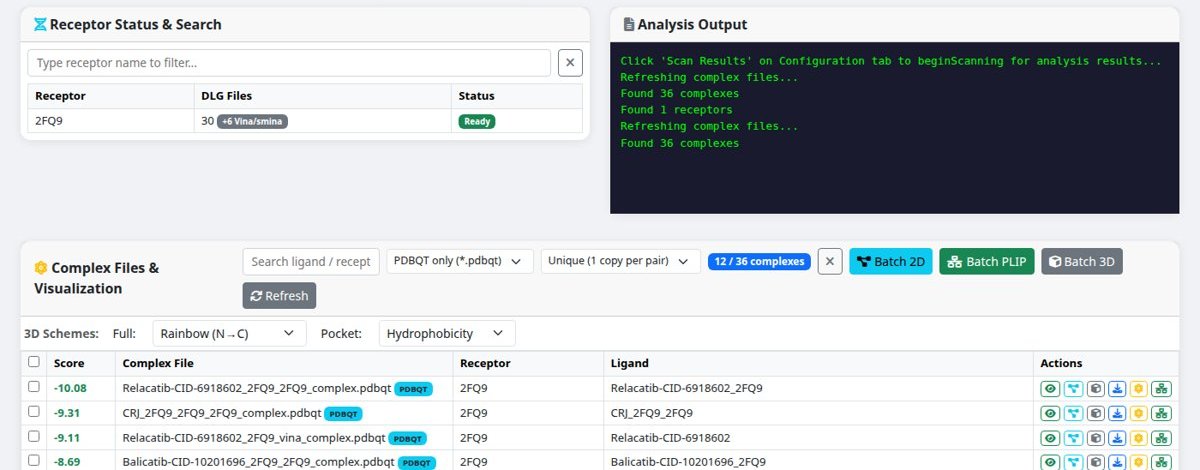
Complexes ranked by score, with batch 2D, 3D and interaction analysis.
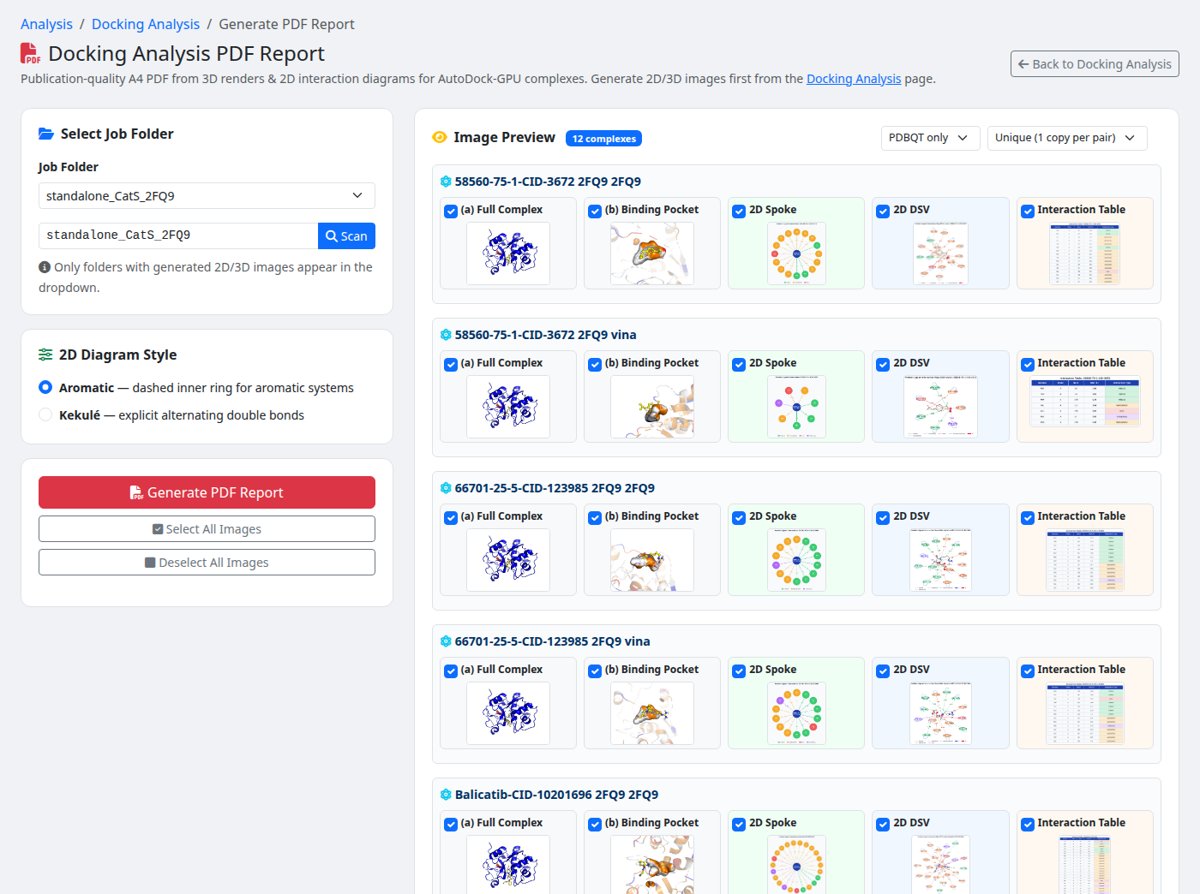
The PDF report: images and interactions of every complex.
Robust ranking
Consensus Docking
- Statistical consensus — AutoDock-GPU at two search depths (SP: 50 runs, XP: 100 runs with 10× longer searches), each in replicates: mean ± SD, weighted consensus score, t-test and effect size, plots
- Multi-engine consensus — AutoDock-GPU, Vina and smina ranked together by mean rank, rank sum or exponential consensus ranking
- Complexes and interactions of the consensus top hits; PDF reports
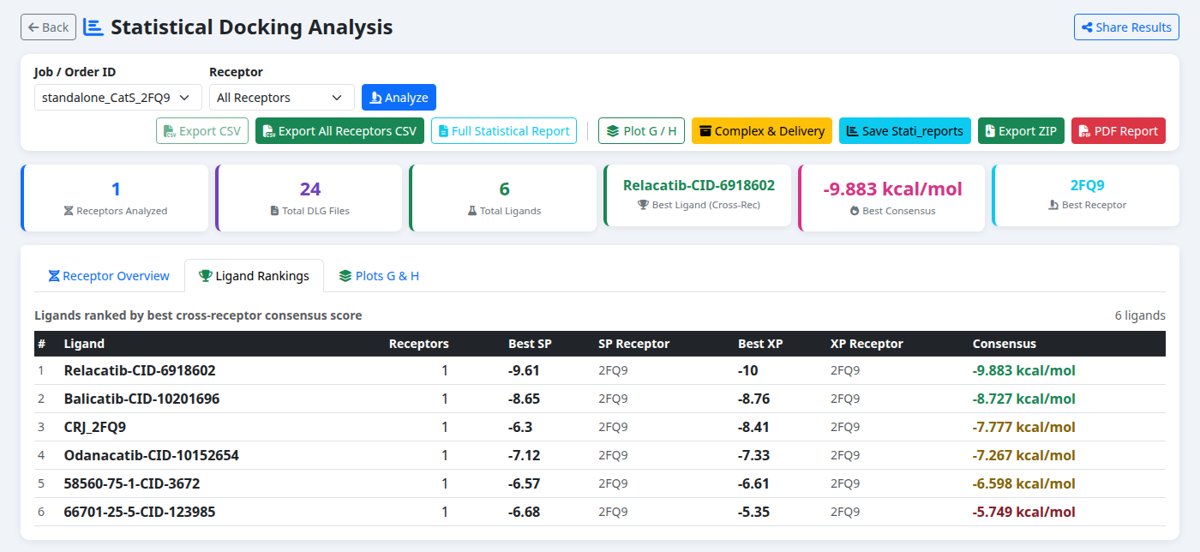
Statistical docking: best SP and XP scores and the consensus per ligand.
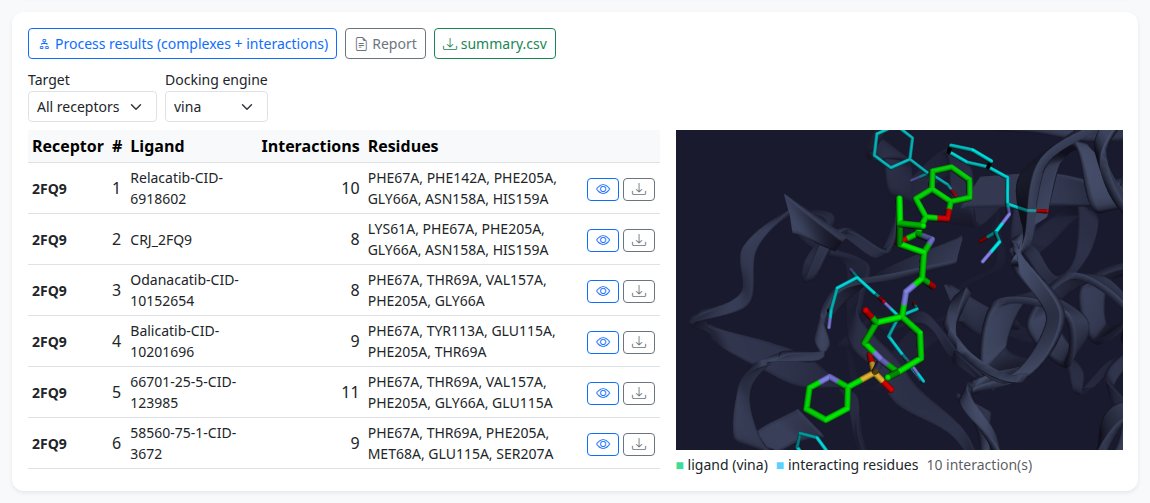
Multi-engine consensus: interactions of each ligand and engine, with a 3D view.
Receptor flexibility
Ensemble Docking
- Several receptor conformations: representatives of an MD simulation, crystal / NMR structures, or an uploaded trajectory
- All conformations superposed, one common box
- Rankings by best, mean, population-weighted and robustness scores; binding modes compared across conformations
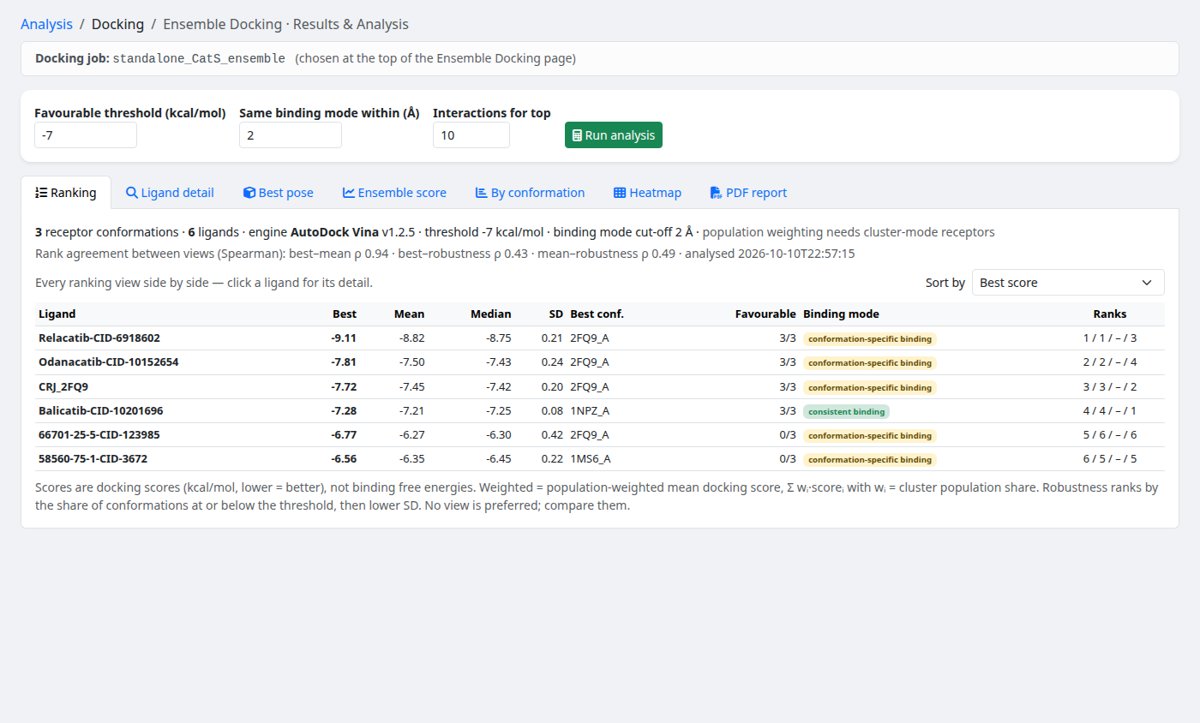
Ranking over three cathepsin S structures.
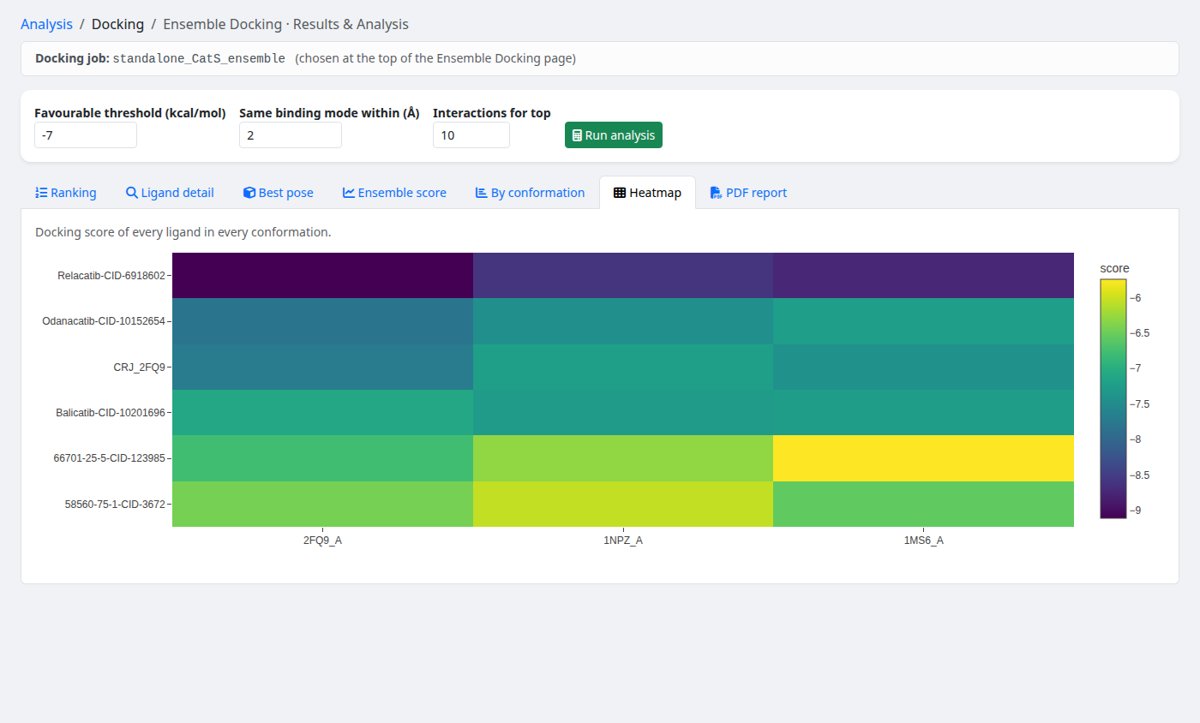
Score of every ligand in every conformation.